

Among the thousands of genes implicated in cancer, few have had as profound an impact as MYC. Unlike many oncogenes that drive a limited number of tumors, MYC acts as a universal amplifier of malignant behavior. It regulates cellular proliferation, metabolism, protein synthesis, mitochondrial function, stemness, angiogenesis, immune evasion, and genomic instability. Estimates suggest that dysregulated MYC activity contributes to 50–70% of all human cancers, making it one of the most pervasive drivers of malignancy. Yet despite more than four decades of intensive research, MYC remains one of oncology’s most elusive therapeutic targets. Its protein structure lacks the deep binding pockets that conventional drugs require, earning MYC the reputation of being the archetypal “undruggable” oncogene.
Recent advances, however, are changing this landscape. Indirect approaches targeting BET proteins, CDK7, Aurora kinases, glutamine metabolism, and synthetic lethal interactions have renewed interest in MYC-directed therapy. Simultaneously, the metabolic theory of cancer offers a complementary framework for understanding MYC. Rather than viewing MYC solely as a mutation that drives uncontrolled proliferation, it can be seen as a master regulator that rewires cellular metabolism toward glycolysis, glutamine addiction, mitochondrial remodeling, and anabolic growth. This perspective integrates MYC naturally into the Five-Axis Metabolic Trap, where metabolic pressure rather than mutation-specific inhibition becomes the central therapeutic strategy.
Introduction
Cancer develops through the accumulation of alterations that confer selective advantages upon individual cells. While mutations such as EGFR, KRAS, BRAF, and PIK3CA activate discrete signaling pathways, MYC occupies a unique position at the convergence of many of these networks. Rather than acting as a simple signaling protein, MYC functions as a master transcription factor controlling the expression of thousands of genes.
Virtually every major oncogenic pathway—including RAS, EGFR, Wnt/β-catenin, PI3K-AKT-mTOR, NOTCH, and Hedgehog—ultimately converges on MYC activation. Once activated, MYC orchestrates a comprehensive transcriptional program that transforms a normal cell into one optimized for continuous growth and division.
Unlike tumor suppressor genes such as TP53 or APC, MYC mutations themselves are relatively uncommon. Instead, MYC activity is increased through gene amplification, chromosomal translocation, increased transcription, enhanced protein stability, or persistent upstream signaling. This explains why MYC activation is observed across nearly every major human malignancy despite relatively infrequent mutations within the MYC gene itself.
Biology of MYC
The MYC family consists of three related genes:
c-MYC (MYC) – the predominant form in human cancers
N-MYC (MYCN) – commonly amplified in neuroblastoma and certain brain tumors
L-MYC (MYCL) – associated primarily with small-cell lung cancer
MYC proteins function by forming heterodimers with MAX, allowing them to bind DNA at specific E-box sequences and regulate transcription. Under normal physiological conditions, MYC expression is tightly controlled and transiently induced following growth factor stimulation. In cancer, this regulation is lost, resulting in persistent MYC activity and constitutive expression of genes involved in cell growth.
MYC regulates nearly 10–15% of the human genome, influencing virtually every aspect of cellular physiology.
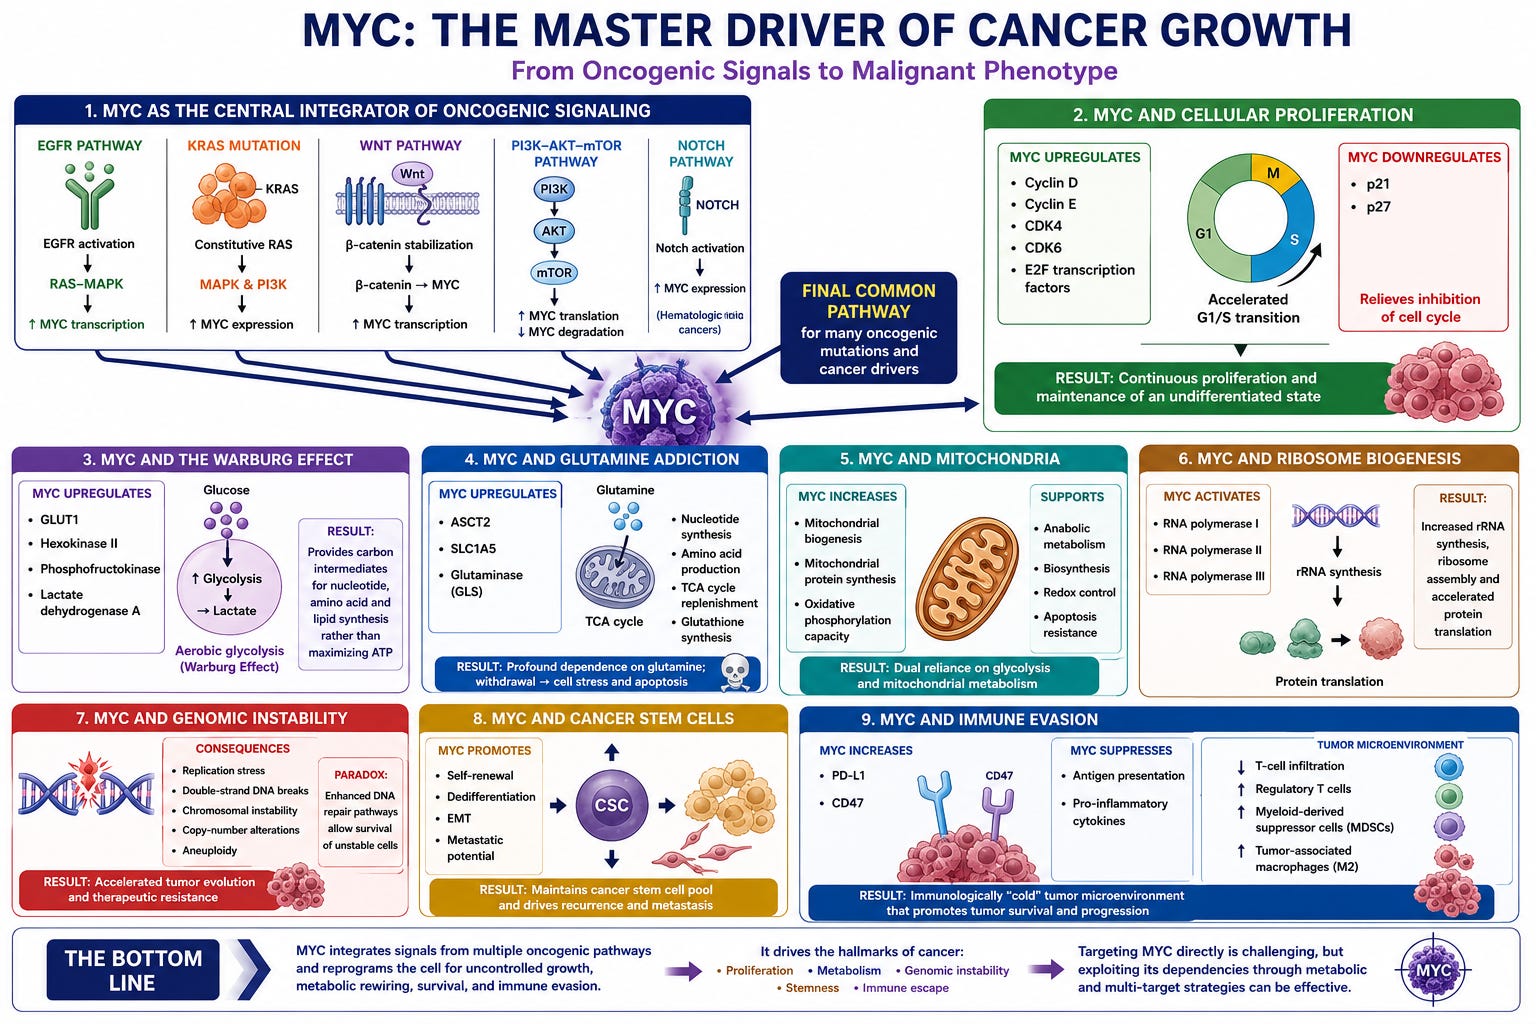
MYC as the Central Integrator of Oncogenic Signaling
One reason MYC has such broad influence is that it sits downstream of multiple oncogenic pathways.
Activation of EGFR stimulates RAS-MAPK signaling, increasing MYC transcription. KRAS mutations sustain MYC expression through MAPK and PI3K signaling. Wnt signaling stabilizes β-catenin, which directly induces MYC transcription. PI3K-AKT-mTOR enhances MYC translation while preventing its degradation. NOTCH signaling similarly increases MYC expression in hematologic malignancies.
Thus, MYC represents the final common pathway through which many oncogenic mutations exert their effects.
MYC and Cellular Proliferation
The hallmark function of MYC is the promotion of cell-cycle progression.
MYC induces expression of:
Cyclin D
Cyclin E
CDK4
CDK6
E2F transcription factors
while simultaneously suppressing cell-cycle inhibitors such as:
p21
p27
The result is accelerated transition through the G1/S checkpoint and continuous proliferation.
Normal tissues carefully balance proliferation with differentiation. MYC disrupts this balance, maintaining cells in an immature, highly proliferative state.
MYC and the Warburg Effect
One of MYC’s most important contributions to cancer biology is metabolic reprogramming.
MYC dramatically increases glucose utilization by inducing expression of:
GLUT1
Hexokinase II
Phosphofructokinase
Lactate dehydrogenase A
This promotes aerobic glycolysis even in the presence of adequate oxygen—a phenomenon first described by Otto Warburg nearly a century ago.
Rather than maximizing ATP production, glycolysis provides rapidly dividing cells with carbon intermediates required for nucleotide, amino acid, and lipid synthesis.
MYC therefore converts metabolism from an energy-generating process into a biosynthetic factory.
MYC and Glutamine Addiction
Glucose alone cannot sustain rapidly dividing cancer cells.
MYC simultaneously creates profound dependence upon glutamine by increasing expression of:
ASCT2
SLC1A5
Glutaminase (GLS)
Glutamine supplies carbon and nitrogen necessary for:
nucleotide synthesis
amino acid production
TCA cycle replenishment
glutathione synthesis
Many MYC-driven tumors become so dependent upon glutamine that withdrawal induces rapid apoptosis.
This metabolic vulnerability has stimulated intense interest in glutaminase inhibitors.
MYC and Mitochondria
Although MYC enhances glycolysis, it does not eliminate mitochondrial function.
Instead, MYC remodels mitochondria to support:
anabolic metabolism
biosynthesis
redox control
apoptosis resistance
MYC increases:
mitochondrial biogenesis
mitochondrial protein synthesis
oxidative phosphorylation capacity
This dual dependence upon glycolysis and mitochondrial metabolism explains why MYC-driven cancers often exhibit remarkable metabolic flexibility.
MYC and Ribosome Biogenesis
Few oncogenes stimulate protein synthesis as powerfully as MYC.
MYC directly activates:
RNA polymerase I
RNA polymerase II
RNA polymerase III
resulting in:
increased ribosomal RNA synthesis
enhanced ribosome assembly
accelerated protein translation
Rapid protein synthesis becomes one of the defining characteristics of MYC-driven cancers.
MYC and Genomic Instability
Excessive MYC activity creates replication stress.
DNA replication becomes increasingly error-prone, leading to:
double-strand DNA breaks
chromosomal instability
copy-number alterations
aneuploidy
Paradoxically, MYC simultaneously promotes DNA repair pathways, allowing genetically unstable cells to survive.
This combination accelerates tumor evolution and therapeutic resistance.
MYC and Cancer Stem Cells
Accumulating evidence suggests MYC contributes to maintenance of cancer stem cells.
MYC promotes:
self-renewal
dedifferentiation
epithelial-mesenchymal transition (EMT)
metastatic potential
MYC expression frequently increases during the acquisition of treatment resistance.
Cancer stem cells rely heavily upon MYC-mediated metabolic adaptation, making MYC particularly relevant to recurrent disease.
MYC and Immune Evasion
MYC influences the tumor microenvironment through several mechanisms.
It increases expression of:
PD-L1
CD47
while suppressing antigen presentation and promoting secretion of immunosuppressive cytokines.
MYC-driven tumors frequently exhibit:
reduced T-cell infiltration
increased regulatory T cells
enhanced myeloid-derived suppressor cells
macrophage polarization toward tumor-promoting phenotypes
Thus MYC helps create an immunologically “cold” tumor microenvironment.
MYC Across Human Cancers
MYC dysregulation is observed in numerous malignancies.
Particularly important examples include:
Burkitt lymphoma
Triple-negative breast cancer
Colorectal cancer
Pancreatic cancer
Hepatocellular carcinoma
Gastric cancer
Ovarian cancer
Non-small-cell lung cancer
Small-cell lung cancer
Medulloblastoma
Neuroblastoma
Multiple myeloma
In Burkitt lymphoma, MYC translocation is the defining molecular abnormality.
In neuroblastoma, amplification of MYCN predicts particularly aggressive disease.
Why MYC Has Been Difficult to Target
Unlike kinases such as EGFR or BRAF, MYC lacks an enzymatic active site.
Instead, MYC functions through protein-protein interactions and DNA binding, creating few opportunities for conventional small-molecule inhibitors.
This structural simplicity has made MYC one of the most challenging therapeutic targets in oncology.
Emerging MYC-Directed Therapies
Although direct MYC inhibitors remain experimental, several indirect strategies have shown promise.
These include:
BET bromodomain inhibitors
CDK7 inhibitors
CDK9 inhibitors
Aurora kinase inhibitors
Glutaminase inhibitors
MDM2 inhibitors
Omomyc (dominant-negative MYC inhibitor)
Synthetic lethal strategies targeting MYC-dependent vulnerabilities
Early clinical trials have demonstrated biological activity, although durable clinical responses remain limited.
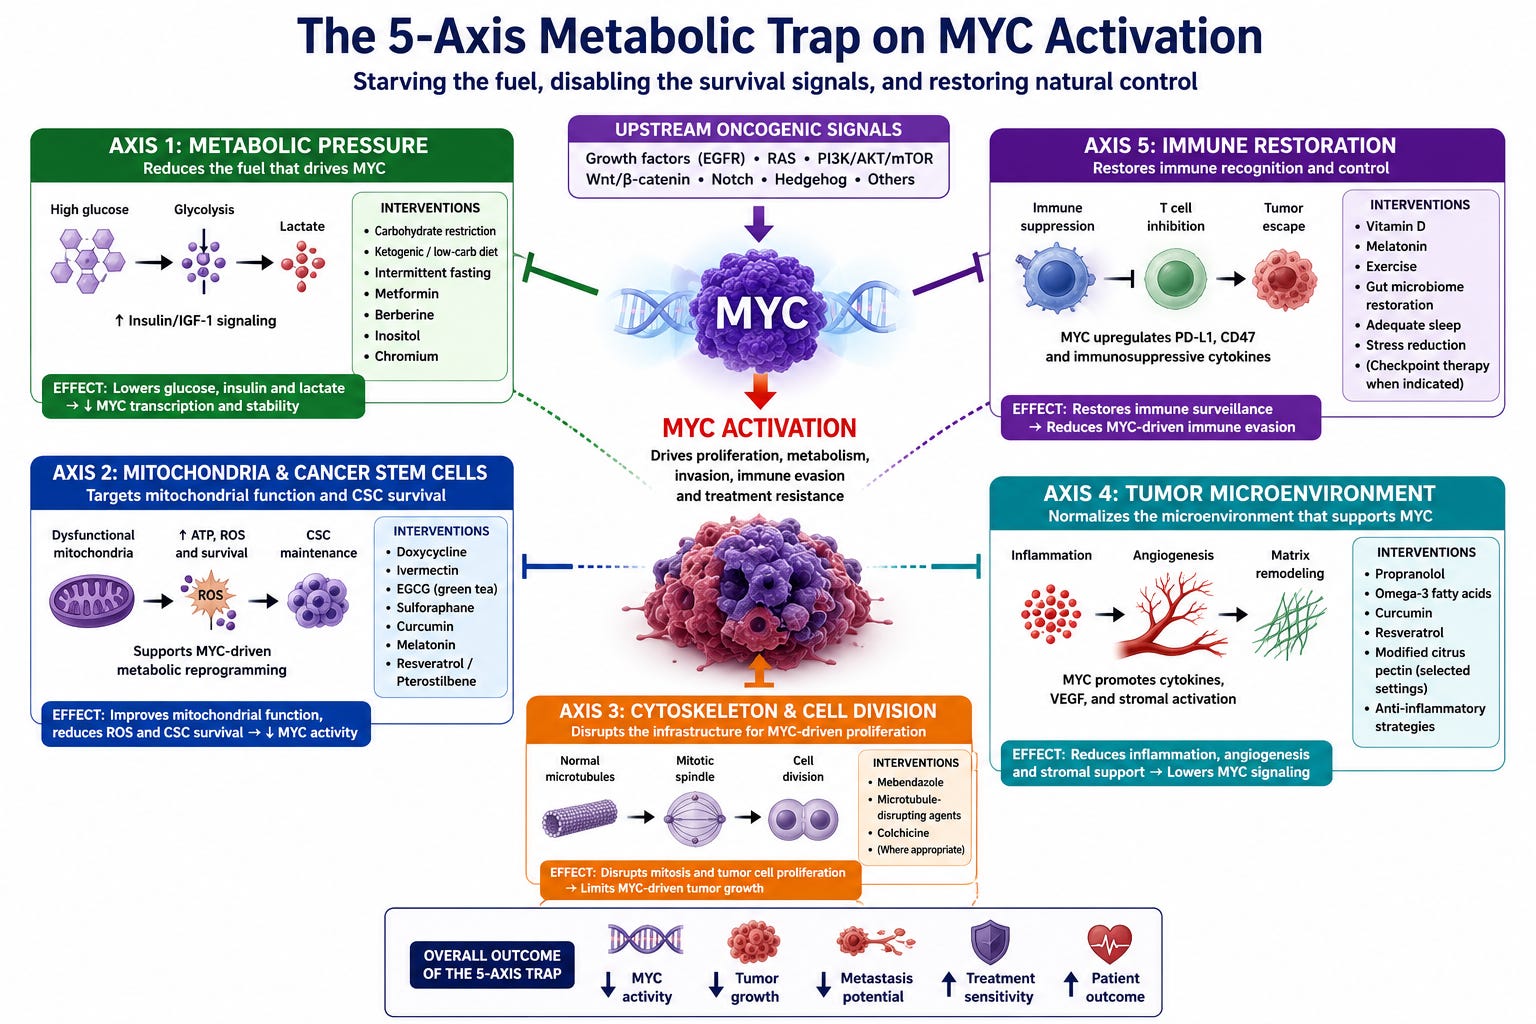
MYC and the Five-Axis Metabolic Trap
Within the Five-Axis framework, MYC is less a single mutation than a master metabolic regulator that simultaneously activates multiple hallmarks of malignant metabolism.
Axis 1 – Metabolic Pressure
MYC markedly increases:
glucose uptake
glycolysis
insulin responsiveness
lactate production
Potential interventions include:
ketogenic or carbohydrate-restricted diets
intermittent fasting
metformin
berberine
Axis 2 – Mitochondria and Cancer Stem Cells
MYC supports mitochondrial remodeling and stem-cell survival.
Potential adjunctive agents include:
doxycycline
ivermectin
EGCG
sulforaphane
curcumin
melatonin
resveratrol or pterostilbene
Axis 3 – Cytoskeleton and Cell Division
Because MYC drives rapid proliferation, dividing cells become increasingly dependent upon intact microtubules.
Potential interventions include:
mebendazole
other microtubule-disrupting agents
Axis 4 – Tumor Microenvironment
MYC promotes angiogenesis, inflammation, and immune suppression.
Potential supportive agents include:
propranolol
omega-3 fatty acids
curcumin
modified citrus pectin (selected settings)
Axis 5 – Immune Restoration
MYC suppresses antitumor immunity through PD-L1 induction and cytokine remodeling.
Supportive strategies may include:
vitamin D optimization
melatonin
regular exercise
restoration of gut microbial diversity
adequate sleep and circadian regulation
Rather than attempting to inhibit MYC directly, the Five-Axis approach seeks to exploit the metabolic dependencies that MYC creates, reducing the tumor’s ability to adapt and survive.
Future Directions
Rapid advances in structural biology, protein degradation technologies, and RNA therapeutics are beginning to make MYC a more tractable target. Direct MYC inhibitors, targeted protein degraders, and synthetic lethal approaches are now entering clinical development. At the same time, increasing recognition of MYC’s central role in metabolic reprogramming suggests that combining emerging MYC-directed therapies with metabolic interventions may prove particularly effective.
Conclusion
MYC occupies a unique position in cancer biology. Rather than functioning as a single oncogenic switch, it acts as the master conductor of malignant growth, integrating signals from diverse pathways into a coordinated program of proliferation, metabolic reprogramming, protein synthesis, genomic instability, stemness, and immune evasion. Dysregulation of MYC contributes to the development and progression of the majority of human cancers, making it one of the most important oncogenes ever discovered.
Despite decades of research, MYC has remained an extraordinarily difficult therapeutic target. Nevertheless, advances in indirect MYC inhibition and synthetic lethal strategies are beginning to challenge the notion that MYC is “undruggable.” Equally important, understanding MYC through the lens of cancer metabolism highlights opportunities to exploit the very dependencies it creates. By combining standard therapies with rational metabolic interventions that target glycolysis, glutamine utilization, mitochondrial function, inflammation, and immune suppression, it may be possible to weaken the metabolic foundation upon which MYC-driven cancers depend. As our understanding of MYC continues to evolve, it is likely to remain at the center of both molecular oncology and metabolism-based therapeutic strategies.

Medical Disclaimer: The discussion of repurposed medications and nutraceuticals in this article is intended to review the scientific literature and does not constitute a recommendation for self-treatment. Decisions regarding the use of off-label therapies should be made in consultation with a qualified healthcare professional familiar with the patient’s medical history and current treatment plan. Please see the full Medical Disclaimer on the introductory page to Marik’s Cancer & Metabolic Healing Playbook.
Yes, fascinating. Short term fasting will likely help.
MYC is MYC.. it is not an abbreviation for anything.
MYC most commonly refers to a family of genes and proteins that control cell growth, division, and metabolism.